


PeptideSpectrumMatching
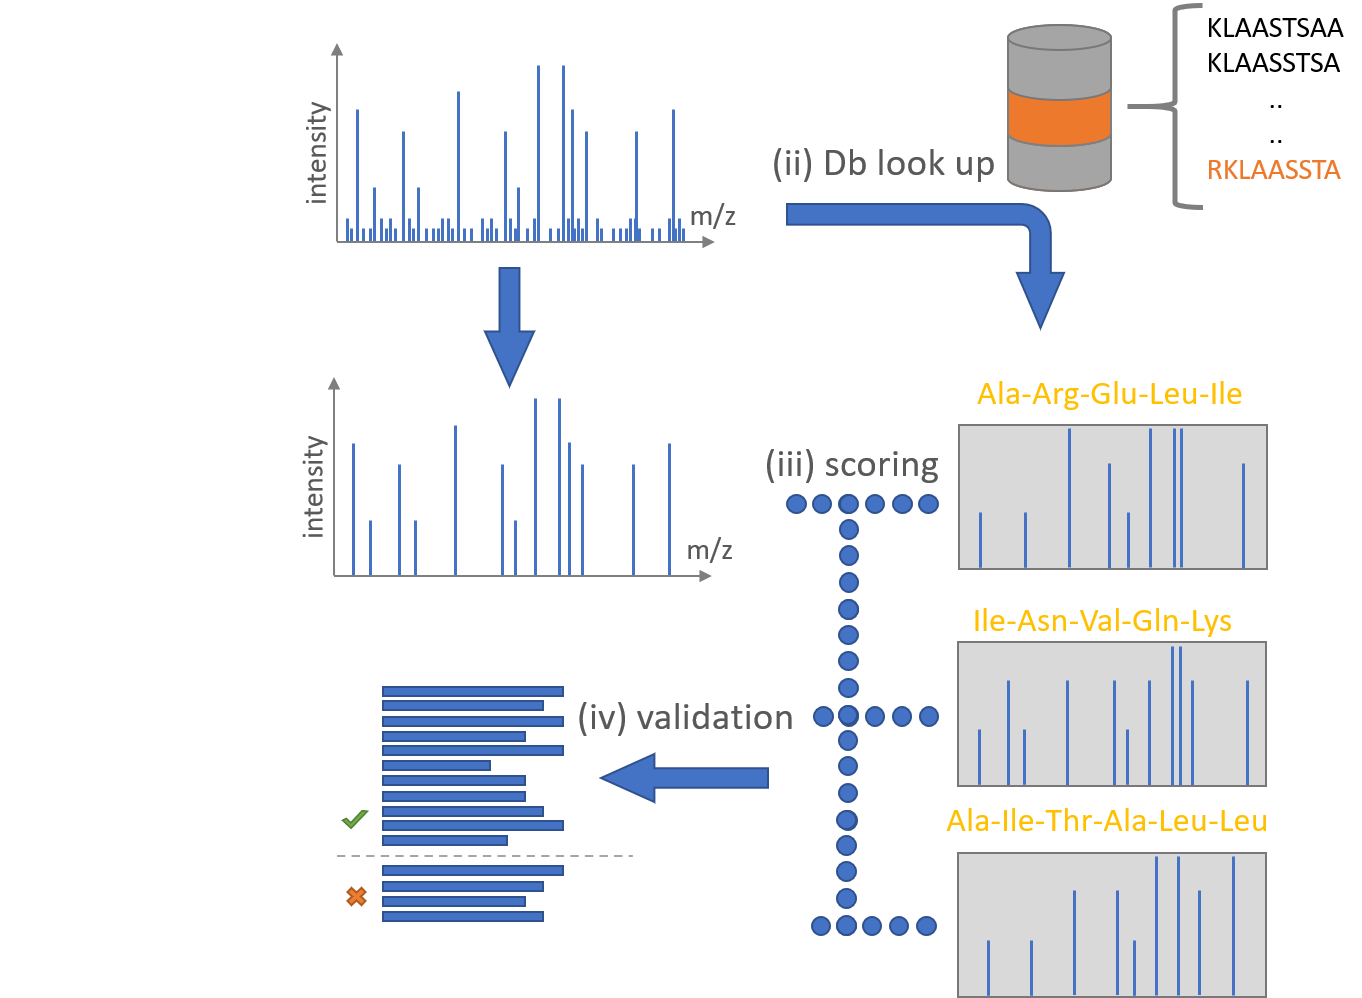
PeptideSpectrumMatching identifies the peptides behind the MS/MS spectra of a run. For each MS2 scan it first determines the charge of the precursor ion from the isotope pattern in the preceding MS1 scan (charge state determination). With the charge, the precursor m/z becomes a peptide mass, and the tool fetches every peptide from the PeptideDB database whose mass lies within LookUpPPM of it. For each candidate it computes the theoretical fragment ions (fragmentation) and compares them with the measured spectrum. Each candidate gets a SEQUEST-like score and an Andromeda-like score. An X!Tandem-like score is computed alongside the Andromeda one. The candidates include reversed decoy peptides, which PSMStatistics later uses to estimate the false discovery rate (what the decoys are for).
Inputs and outputs
Flag |
Meaning |
Comes from |
|---|---|---|
|
one or more |
|
|
the SQLite peptide database |
|
|
the output directory, created when missing |
|
|
the parameter file in JSON |
this page |
|
number of runs scored at the same time, default 1 |
All four flags are mandatory. The reader opens files by extension and knows .mzlite and .mzML only, so an mzML file has to carry the extension in exactly that spelling. The tool stops when the database file does not exist.
The tool writes one <run>.psm per input into the output directory. It is a tab separated table with a header. Each row is one candidate peptide for one spectrum with its charge, precursor m/z, theoretical mass, the three scores with their delta values to the next candidates, the peptide sequence and a Label of 1 for a target peptide and -1 for a decoy.
The file is opened in append mode, so running the tool twice into the same directory adds a second header and a second set of rows. Delete the old file first.
The .psm is read by PSMStatistics. The output directory also receives PeptideSpectrumMatching_log.txt and one <run>_log.txt per input.
Parameters
Parameter |
Default |
Meaning |
|---|---|---|
ChargeStateDeterminationParams |
see below |
Settings of the charge state determination. |
LookUpPPM |
30.0 |
Mass window, in ppm around the precursor mass, from which candidate peptides are fetched. |
nTerminalSeries |
NTerminalSeries.B |
Fragment ion series counted from the N terminus. Keep B, the scoring builds the theoretical spectra from the b and y series. |
cTerminalSeries |
CTerminalSeries.Y |
Fragment ion series counted from the C terminus. Keep Y. |
Andromeda |
{ PMinPMax = 4, 10; MatchingIonTolerancePPM = 100.0 } |
Settings of the Andromeda-like and X!Tandem-like scoring. |
ChargeStateDeterminationParams is a BioFSharp.Mz ChargeState.ChargeDetermParams record.
Parameter |
Default |
Meaning |
|---|---|---|
ExpectedMinimalCharge |
2 |
Lowest precursor charge that is tested. |
ExpectedMaximumCharge |
5 |
Highest precursor charge that is tested. |
Width |
1.1 |
Width of the m/z window around the precursor in which the MS1 isotope pattern is examined. |
MinIntensity |
0.15 |
Minimum relative intensity of a peak to count as part of the isotope pattern. |
DeltaMinIntensity |
0.3 |
Minimum relative intensity difference used when comparing neighbouring peaks of the pattern. |
NrOfRndSpectra |
10000 |
Number of random spectra used to estimate how likely a matching pattern arises by chance. |
Andromeda is a ProteomIQon.Domain.AndromedaParams record.
Parameter |
Default |
Meaning |
|---|---|---|
PMinPMax |
4, 10 |
Lowest and highest number of most intense peaks kept per 100 Da window. Every count in between is tried and the best score is kept. |
MatchingIonTolerancePPM |
100.0 |
Tolerance in ppm for matching a theoretical fragment to a measured peak. |
The default file is peptideSpectrumMatchingParams.json.
Writing a parameter file
open BioFSharp.Mz
open ProteomIQon

let chargeDetermParams : ChargeState.ChargeDetermParams =
    {
        ExpectedMinimalCharge = 2
        ExpectedMaximumCharge = 5
        Width                 = 1.1
        MinIntensity          = 0.15
        DeltaMinIntensity     = 0.3
        NrOfRndSpectra        = 10000
    }

let andromedaParams : Domain.AndromedaParams =
    {
        PMinPMax                = 4, 10
        MatchingIonTolerancePPM = 100.0
    }

let peptideSpectrumMatchingParams : Dto.PeptideSpectrumMatchingParams =
    {
        ChargeStateDeterminationParams = chargeDetermParams
        LookUpPPM                      = 30.0
        nTerminalSeries                = NTerminalSeries.B
        cTerminalSeries                = CTerminalSeries.Y
        Andromeda                      = andromedaParams
    }

// Replace the temp folder with your project folder.
let outputPath = System.IO.Path.Combine(System.IO.Path.GetTempPath(), "peptideSpectrumMatchingParams.json")

Json.serializeAndWrite outputPath peptideSpectrumMatchingParams
Running the tool
Install the tool with dotnet tool install --global ProteomIQon.PeptideSpectrumMatching, then score one run:
|
Several runs, three of them scored at the same time:
|
All flags:
|
module PeptideSpectrumMatchingParams from ProteomIQon.Dto
--------------------
type PeptideSpectrumMatchingParams = { ChargeStateDeterminationParams: ChargeDetermParams LookUpPPM: float nTerminalSeries: NTerminalSeries cTerminalSeries: CTerminalSeries Andromeda: AndromedaParams } member Equals: PeptideSpectrumMatchingParams * IEqualityComparer -> bool
module NTerminalSeries from ProteomIQon.Common
--------------------
type NTerminalSeries = | A | B | C | AB | AC | BC | ABC member Equals: NTerminalSeries * IEqualityComparer -> bool member IsA: bool member IsAB: bool member IsABC: bool member IsAC: bool member IsB: bool member IsBC: bool member IsC: bool
module CTerminalSeries from ProteomIQon.Common
--------------------
type CTerminalSeries = | X | Y | Z | XY | XZ | YZ | XYZ member Equals: CTerminalSeries * IEqualityComparer -> bool member IsX: bool member IsXY: bool member IsXYZ: bool member IsXZ: bool member IsY: bool member IsYZ: bool member IsZ: bool
<summary>Performs operations on <see cref="T:System.String" /> instances that contain file or directory path information. These operations are performed in a cross-platform manner.</summary>
System.IO.Path.Combine([<System.ParamArray>] paths: string array) : string
System.IO.Path.Combine(path1: string, path2: string) : string
System.IO.Path.Combine(path1: string, path2: string, path3: string) : string
System.IO.Path.Combine(path1: string, path2: string, path3: string, path4: string) : string